FDA QMSR Consulting Services

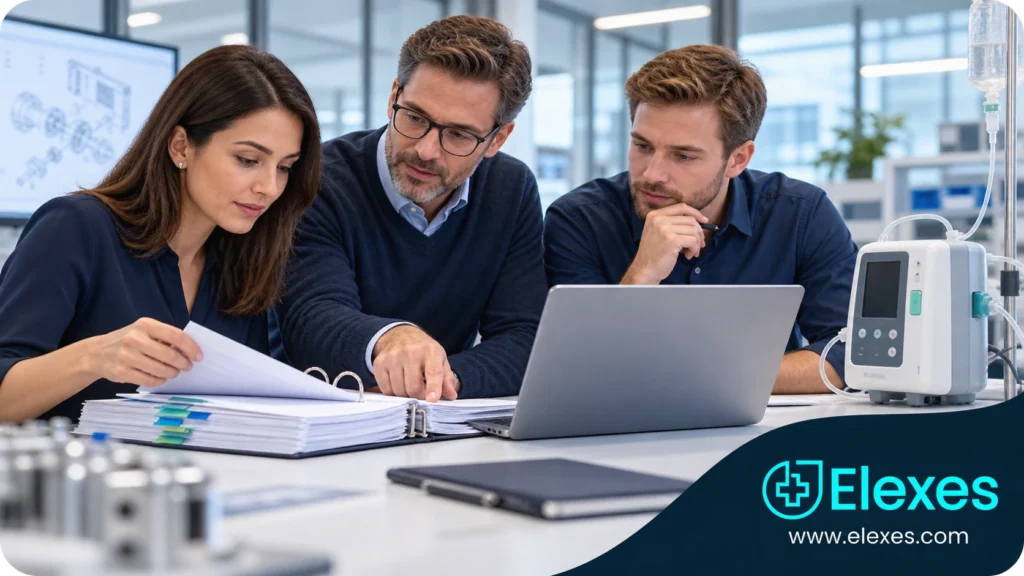
Medical device quality and regulatory professionals reviewing QMSR procedures and records beside an infusion pump.

Make your quality system work for today’s FDA requirements.

Your procedures may look complete. But can your team connect a complaint, a supplier change or a release decision to the records FDA expects?

Elexes helps medical device companies assess, implement and remediate their systems for 21 CFR 820 QMSR, with clear responsibilities and evidence that reflects daily work.

We support startups building their first QMS, growing manufacturers entering the US market and established companies addressing gaps. Founders, executives and RA, QA, engineering, operations, manufacturing, product and compliance teams gain a practical plan.

What Does 21 CFR 820 QMSR Require Now?

The FDA Quality Management System Regulation became applicable as from February 2, 2026. The regulation integrates terms contained in ISO 13485:2016 and ISO 9000:2015 Section 3 while incorporating the FDA regulatory standards.

Compliance means an implemented QMS, appropriate records and conformity with other applicable FDA obligations. ISO 13485 certification is not required by QMSR and does not replace FDA compliance or exempt a manufacturer from inspection.

Our ISO 13485 consulting services cover the common QMS base; our US requirements consulting services take care of the rest.

Where Elexes Focuses Your QMSR Assessment

Scope, Responsibilities and Risk-Based Processes

Identify the operations and equipment that qualify as 21 CFR Part 820. Map the requirements to the appropriate owner and evidence. Device manufacturers cover those that produce specifications and companies involved in manufacturing, sterilizing, re-labeling, or re-packaging in a contract manner.

Component suppliers, importers, and distributors must have role-specific evaluations because not all companies have the same Part 820 requirements. Review the exemption of devices based on classifications. Some devices that are exempted still need to maintain complaint files and records.

We connect management oversight, competence, risk management, suppliers, production and improvement. Applicable design and development requirements are assessed using section 820.10(c), including Class II, Class III and specified Class I devices.

FDA-Specific Records and Reporting Interfaces

ISO 13485 is supplemented with Section 820.35 that contains complaints, servicing and UDI records. We examine form requirements, information that should be included, investigation decisions and relationships between records.

A complaint workflow must also support applicable Medical Device Reporting decisions. Other interfaces include corrections and removals, UDI and device tracking where applicable. These obligations retain their own regulatory requirements. 

Focused complaint handling support can address intake, investigation, escalation and documentation gaps.

Labeling, Packaging and Release Evidence

Section 820.45 includes additional requirements for labeling and packaging controls, accuracy verifications, documentation for labeling release, and precautions to avoid mix-ups. We verify the process by which authorized information is transferred to the production floor and inspections are documented.

A revised label must be submitted to the medical device file, instruction of work, inspection process, and release records. Our production and service controls assistance can help establish such links.

Evidence for Current FDA Inspections

FDA now uses Compliance Program 7382.850. The inspection approach of FDA differs from MDSAP audit. FDA may consider the results of management review, quality audit, and supplier audit, and also documents generated prior to the QMSR effective date.

Recent activities are reviewed in order to verify compliance between procedures and practices. In order to prepare well, our medical device internal audit services will be able to pinpoint deficiencies.

Medical device quality and manufacturing professionals checking packaging labels against controlled records.

How the Engagement Works

Assess: Review your device scope, manufacturing model, existing QMS and representative records. Identify applicable requirements and prioritize gaps.

Implement: Decide on modifications to documentation, FDA record documentation process, training, and assignments. Consider available evidence where appropriate; identify all gaps.

Verify: Check completed actions against recent work and confirm that staff can retrieve records and explain decisions.

Deliverables may include gap analysis, requirements to evidence matrix, priority list of actions, procedures or forms to agree on, and closure log. Work can be performed remotely including secured document reviews and workshops; on-site activities are determined based on need.

RA interprets regulatory obligations, QA coordinates system controls, and engineering, purchasing, manufacturing and service owners implement changes. Leadership provides resources and accountability. Submission preparation, laboratory testing and validation execution are separately scoped.

Parul Chansoria, founder of Elexes, on connecting FDA-specific records with daily quality system work.

Check your scope, FDA-specific records, labeling controls and inspection evidence. Use the one-page checklist to identify gaps and assign follow-up.

FAQs

Frequently Asked Questions!

No. Assess FDA-specific provisions and other applicable US requirements alongside the shared QMS framework.

No. Evaluate existing evidence against current requirements and remediate identified gaps. Preserve accurate history; do not backdate records.

FDA does not issue ISO 13485 certificates through QMSR. Its inspections assess regulatory compliance.

Define applicable responsibilities and controls for outsourced processes. The arrangement does not automatically remove your manufacturer’s obligations.

Close the Gaps That Matter to Your US Operations

Start with a clear view of your current system, the evidence it produces and the actions needed next. Our QMS standards and compliance services connect this work with your wider quality strategy.

A Trusted Team of Consultants

Elexes offers a globally recognized ISO 13485 certification consultant who is renowned to enhancing your business credibility in the medical devices industry.

Local Language
Auditor

Our presence across the globe ensures audits are conducted by local-language auditors, facilitating clear communication and understanding.
Why Choose Elexes?

Expertise

Our experts provide guidance and consultants to help you adhere to necessary regulations, ensuring your products meet all legal requirements and is compliant to all necessary standards.

Local Regulatory Certification

At Elexes, we offer to assist in securing additional local certifications, enabling you to manufacture and sell medical devices in various markets across the globe.

Confidentiality

We offer complete confidentiality to all our clients which makes us one of the most trusted regulatory and quality compliance consulting.

Associated Regulatory Authorities.

img
We’ve More
Than 256+ Global Clients
The Best RA & QA Consultants
Clients Testimonials

Amazing Feedback About Our Services

Popular Clients

We’ve 200+ Global Premium Clients

Saneso
Niramai
Apex dental sensors
Neurasignal
SS inovations
Amaryllis
Inclode
iMedisync
DJOGlobal logo cmyk_TM
Summus Laser Logo
Looking For Regulatory Assistance?

Let's Talk How We Can Help You?

Please enable JavaScript in your browser to complete this form.
Name
img
Please enable JavaScript in your browser to complete this form.
Name
img
Please enable JavaScript in your browser to complete this form.
Name
img
Please enable JavaScript in your browser to complete this form.
Name
img
Please enable JavaScript in your browser to complete this form.
Name
img